Non-hydroxamate LpxC inhibitors
A Vernalis and Taisho collaboration designed around the hydroxamate zinc-binding group, delivering an antibacterial lead active against Pseudomonas aeruginosa.
The Challenge
UDP-3-O-acyl-N-acetylglucosamine deacetylase (LpxC) is a zinc metalloenzyme that catalyses the first committed step in the biosynthesis of Lipid A, an essential component of the cell envelope of Gram-negative bacteria. The most advanced disclosed LpxC inhibitors showing antibacterial activity coordinate zinc through a hydroxamate moiety, raising concerns about binding to other metalloenzymes.
The Science
A collaboration between Vernalis Research and Taisho Pharmaceuticals used fragment and structure-based methods for the discovery and optimisation of two series of compounds with differing modes of zinc coordination.
One series was evolved from a fragment in which a glycine moiety complexes zinc. It achieved low nanomolar potency in an enzyme functional assay, but poor antibacterial activity in cell cultures.
A second series was based on a fragment that coordinated zinc through an imidazole moiety. Structure-guided design led to a 2-(1S-hydroxyethyl)-imidazole derivative, compound 43, exhibiting low nanomolar inhibition of LpxC and a minimum inhibitory concentration (MIC) of 4 ug/mL against Pseudomonas aeruginosa, which is little affected by the presence of albumin.
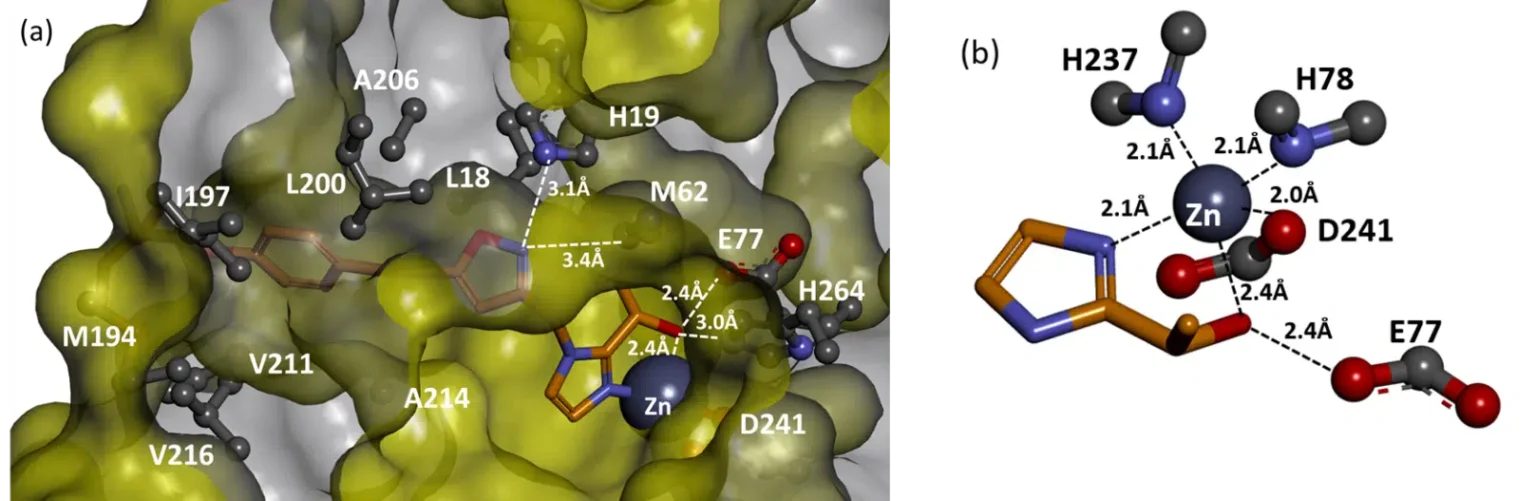
Crystal structure of compound 43 bound to the active site of PaLpxC, showing zinc coordination to the ligand and protein side chains.
Key Takeaways
- Two compound series developed with differing zinc coordination, avoiding the hydroxamate liability
- Compound 43 combines low nanomolar LpxC inhibition with an MIC of 4 ug/mL against P. aeruginosa
- Activity little affected by albumin — a practical developability signal
- The only anti-infective programme in the Vernalis case study portfolio
Service Areas
Work with us
Vernalis has collaborated with a wide range of partners to enable some of the most challenging therapeutic targets. Present and past collaborators include Servier, Pierre Fabre, Genentech and Lundbeck. Vernalis can support programs with DEL and fragment-based technologies through to establishing full drug discovery programmes for our partners.
Our track record is built on innovation and success in enabling drug discovery and identifying clinical candidates for a wide variety of therapeutic targets.
Research Director and Joint Site Lead
James Murray PhD
Similar Case Studies
Explore how our team overcomes challenges through case studies showcasing our success stories.
-
Hsp90 oral backup pre-clinical candidate
Fragment-based and in silico hits were combined into a novel series of Hsp90 inhibitors, delivering tumour growth regression on oral dosing in a BT474 xenograft model.
Outcome Oral pre-clinical candidate — the follow-on to the intravenous phase II compound. -
DYRK1A and DYRK1B selective inhibitors
A Vernalis and Servier NMR fragment screen delivered two series of potent, selective DYRK1A inhibitors, including the brain-penetrant tool compound 34.
Outcome Two optimised inhibitor series and a selective, brain-penetrant tool compound. -
LRRK2 crystallographic surrogate
Vernalis engineered crystallographic surrogates of LRRK2 from Chk1 mutants, then used them to design a potent, brain-penetrant inhibitor with confirmed target engagement.
Outcome Target enablement — a crystallisable system, and a brain-penetrant inhibitor designed against it. -
PDHK — application of off-rate screening
SPR off-rate screening of unpurified reaction products drove rapid hit-to-lead progression to selective, sub-100 nM pan-isoform PDHK inhibitors.
Outcome Proprietary off-rate screening validated on a live programme, delivering sub-100 nM leads. -
Hsp90 phase II candidate
Fragment, virtual and medium-throughput screening produced NVP-AUY922, a well-tolerated Hsp90 inhibitor that progressed into global phase II trials.
Timeline Phase I entry 2007 to clinical proof of concept, April 2011.Outcome Clinical proof of concept, April 2011. -
Bcl-2 phase I clinical candidate
Structure-guided design produced S55746, an orally bioavailable and Bcl-2-selective inhibitor that spares platelets and reached phase I.
Outcome Phase I clinical candidate with a clean selectivity and tolerability profile. -
Mcl-1 in vivo active inhibitor
An NMR fragment screen and structure-guided design produced S63845, a specific, high-affinity Mcl-1 inhibitor with single-agent anti-tumour activity in vivo.
Outcome In vivo active inhibitor of a target long considered difficult to drug -
Chk1 pre-clinical application
Structure-guided fragment elaboration delivered V158411, a potent Chk1/Chk2 inhibitor that potentiates irinotecan without added systemic toxicity.
Outcome Pre-clinical candidate demonstrating combination efficacy without added toxicity.
Start your own success story
We’d love to hear from you! Head to our contact page to get started.